
The denecimig FDA review has been extended because remediation work is still under way at a manufacturing facility, Novo Nordisk said on 2 October 2026. The US Food and Drug Administration (FDA) has not identified deficiencies in the clinical efficacy or safety data for the experimental hemophilia A treatment, according to the company announcement, and has not communicated a new decision date. Novo now expects a US launch in the first half of 2027, subject to approval, and says its 2026 financial outlook is unchanged.
Key Takeaways
- The FDA has extended its review of denecimig, Novo Nordisk’s hemophilia A therapy, because manufacturing facility remediation is still ongoing.
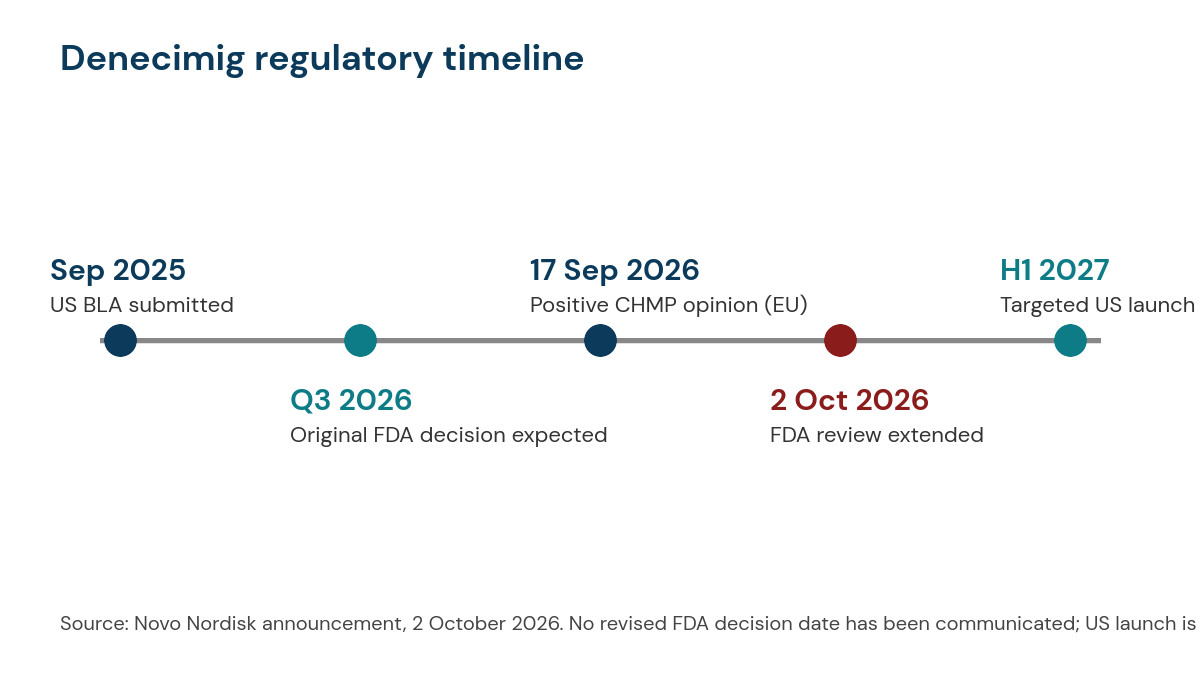
- A decision had been expected in the third quarter of 2026, after the biologics license application (BLA) was submitted in September 2025. No revised date has been given.
- Novo says the FDA has not identified deficiencies related to clinical efficacy or safety data.
- The company now targets a US launch in the first half of 2027, pending approval, and keeps its 2026 outlook unchanged.
- In Europe, the CHMP adopted a positive opinion on 17 September 2026.
What Happened in the Denecimig FDA Review?
Novo Nordisk said the FDA has extended its review of the denecimig BLA while remediation activities continue at a manufacturing site. The issue arose after a pre-license inspection of the facility, and the company says it is working with the agency on the remaining items. Novo did not name the facility or give its location in the announcement we reviewed.
Mike Doustdar, Novo Nordisk president and chief executive, said: “Novo is already in the process of addressing the FDA’s requests as efficiently as possible and continuing to work closely with the Agency toward bringing denecimig to patients in the US.” The statement is reported by Securities.io.
Denecimig FDA Review at a Glance
| Item | Detail |
|---|---|
| Drug | Denecimig (Novo Nordisk), FVIIIa mimetic bispecific antibody |
| Indication sought | Prophylaxis in hemophilia A, with or without inhibitors, adults and children |
| BLA submitted | September 2025 |
| Original US decision expected | Third quarter of 2026 |
| Reason for extension | Manufacturing facility remediation ongoing |
| Efficacy or safety deficiencies | None identified by FDA, per Novo |
| Revised FDA decision date | Not communicated |
| Targeted US launch | First half of 2027, pending approval |
| Europe | Positive CHMP opinion, 17 September 2026 |
| Dosing | Weekly, every two weeks or monthly, subcutaneous |
What Is Denecimig?
Denecimig is an experimental bispecific antibody for hemophilia A, a bleeding disorder caused by a lack of working clotting factor VIII. According to Novo, it is a factor VIIIa mimetic: it brings together two clotting proteins, factor IXa and factor X, to do the job that missing factor VIII normally does. It is given as an injection under the skin.
Novo is seeking approval for routine prophylaxis, meaning regular treatment to prevent bleeds, in adults and children with hemophilia A, with or without inhibitors. Inhibitors are antibodies that can make factor VIII replacement less effective. The company says patients could choose between dosing once a month, every two weeks or weekly. Denecimig would join a field in which Roche’s Hemlibra is the established bispecific therapy; Stocktwits cites one estimate that Hemlibra generated about $5.8 billion in sales last year and an analyst view that denecimig could reach peak annual sales near $2 billion. Those are third-party estimates, not company forecasts.
The Clinical Data Behind Denecimig
The filing is based on the FRONTIER clinical programme, including the FRONTIER2, FRONTIER3 and FRONTIER4 trials. Securities.io reports that annualised bleeding rates were generally below one in the phase 3 population, and that a substantial share of participants had no treated bleeds. A further study, FRONTIER5, reportedly found no new safety signals in people switching from Hemlibra, and showed a preference for the device. These results come from Novo and have not been independently reviewed by us.
Why the Review Was Extended: Manufacturing, Not Clinical Data
Drug approvals depend on both clinical evidence and the quality of the facility that makes the product. For biologic medicines, regulators inspect manufacturing sites before approval because even a well-performing drug cannot be supplied if the plant does not meet standards. In this case, Novo says the open points relate to the facility and not to clinical efficacy or safety, and that the issue does not affect its other marketed products.
That distinction matters for readers: manufacturing issues can often be fixed, but the timing is uncertain because the agency must be satisfied before it acts. Facility quality is a theme across the industry as companies build new capacity, as in the Bayer plan for a pharmaceutical plant in Ohio.
What Happens Next
- FDA decision: no new date has been communicated. Novo targets a US launch in the first half of 2027, pending approval.
- Europe: after the positive CHMP opinion on 17 September 2026, the European Commission decision is the next step, and one report indicates launches in select countries from the fourth quarter of 2026.
- Financials: Novo says there is no impact on its 2026 outlook.
What It Means for Patients and the Market
For people with hemophilia A, the delay means a new treatment option will reach the United States later than hoped, although existing therapies remain available. The World Federation of Hemophilia provides information on treatment access worldwide.
For Novo, the setback comes while the company is also investing in new research, as in its Novo Nordisk and Orbis Medicines drug-discovery deal and its Novo Nordisk partnership with Anthropic. Regulators have also been busy with rare disease decisions, such as the FDA approval of Fayuvi for Sanfilippo syndrome type A and the FDA approval of Mirum Atebrioz.
Our Assessment
In our assessment, the denecimig FDA review extension looks like a delay rather than a rejection: the stated issue is manufacturing, the agency has not raised clinical concerns, and Novo has held its outlook. The key unknowns are how long remediation will take and whether the FDA needs to re-inspect. Investors and clinicians should watch for the next regulatory update before relying on the first-half 2027 target.
Frequently Asked Questions
Why did the FDA extend the denecimig review?
Novo says remediation work is still ongoing at a manufacturing facility identified in a pre-license inspection.
Did the FDA raise safety or efficacy concerns?
No. Novo says the agency has not identified deficiencies related to clinical efficacy or safety data.
When will denecimig be available in the US?
Novo now expects a US launch in the first half of 2027, subject to FDA approval.
What is denecimig used for?
It is being developed as routine prophylaxis for hemophilia A, in adults and children with or without inhibitors.
Has denecimig been approved anywhere?
The European CHMP adopted a positive opinion on 17 September 2026. A formal European Commission decision follows that step.
How we reported this: information comes from the Novo Nordisk announcement dated 2 October 2026 and trade coverage by Securities.io and Stocktwits. Clinical and sales figures are company-reported or third-party estimates and are not independently verified. Last updated 3 October 2026. This article is for information only and is not medical or investment advice.
[…] FDA Cancels Cipla Pune Warehouse Drug Licences FDA Extends Review of Novo Denecimig Over Facility Issues Pennsylvania Measles Outbreak Nears 1,000 With 977 Cases Bayer Ohio Plant: $2.2 Billion Pharma […]